 Polynomial scaling localized active space unitary selective coupled cluster singles and doubles
Journal of Chemical Theory and Computation (2025)
Researchers have developed a more efficient preselection step for quantum algorithms to help quantum computers simulate complex molecules.
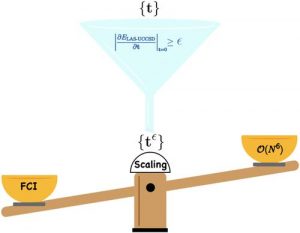
The method focuses only on the most important parts of the molecule and selects the most relevant quantum interactions using a smart filtering system.
The team tested this method on several molecular systems, including chains of carbon atoms and a chromium-based molecule, to check how accurate and practical the results are.
Their approach reduces memory demands for selection and helps estimate the quantum resource requirements for simulating larger, more realistic chemical systems in the future — something current quantum hardware can’t yet handle.
Shreya Verma, Ruhee D’Cunha, Abhishek Mitra, Matthew Hermes, Stephen Gray, Matt Otten and Laura Gagliardi conducted this research, working at the University of Chicago, the University of Wisconsin–Madison and Argonne National Laboratory.